
Process performance qualification (PPQ) is the second element of Stage 2 of process validation in which a manufacturer runs commercial batches, with qualified equipment, trained staff, real components and the commercial process, under an approved protocol to show the process is reproducible at commercial scale. FDA expects PPQ to be complete before commercial distribution of a drug.
This guide adds what PPQ explainers leave out: what FDA investigators wrote when a facility had no evidence that its process performance was validated. The counts come from the Atlas Form 483 citation dataset (pulled October 21, 2026), covering 774 drug inspections that cited 21 CFR Part 211 and closed between January 7, 2025 and September 3, 2026, joined to FDA's inspection classifications.
Key facts:
- FDA's guidance "Process Validation: General Principles and Practices" (January 2011, Revision 1) splits Stage 2 into two elements: qualification of the facility, utilities and equipment, then PPQ.
- The 2011 guidance names no number of PPQ batches; it asks for enough samples to give statistical confidence of quality within a batch and between batches.
- EU GMP Annex 15 (in operation since October 1, 2015), paragraph 5.20, calls a minimum of three consecutive batches made under routine conditions "generally considered acceptable", and paragraph 5.19 still requires each manufacturer to justify its number.
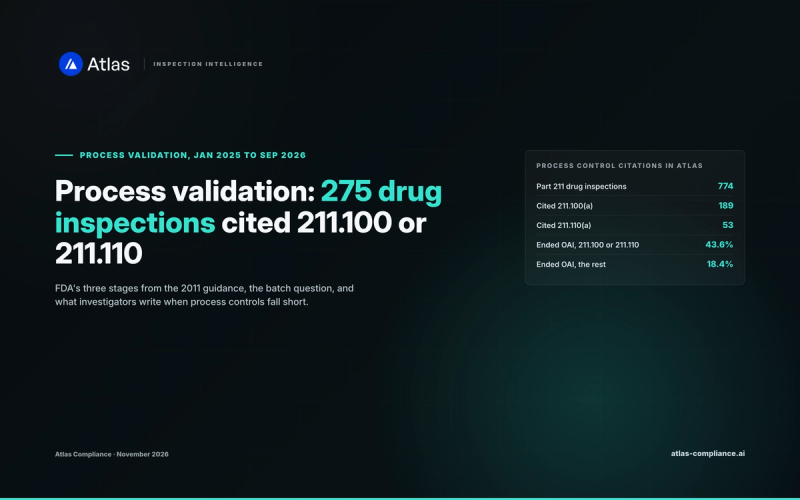
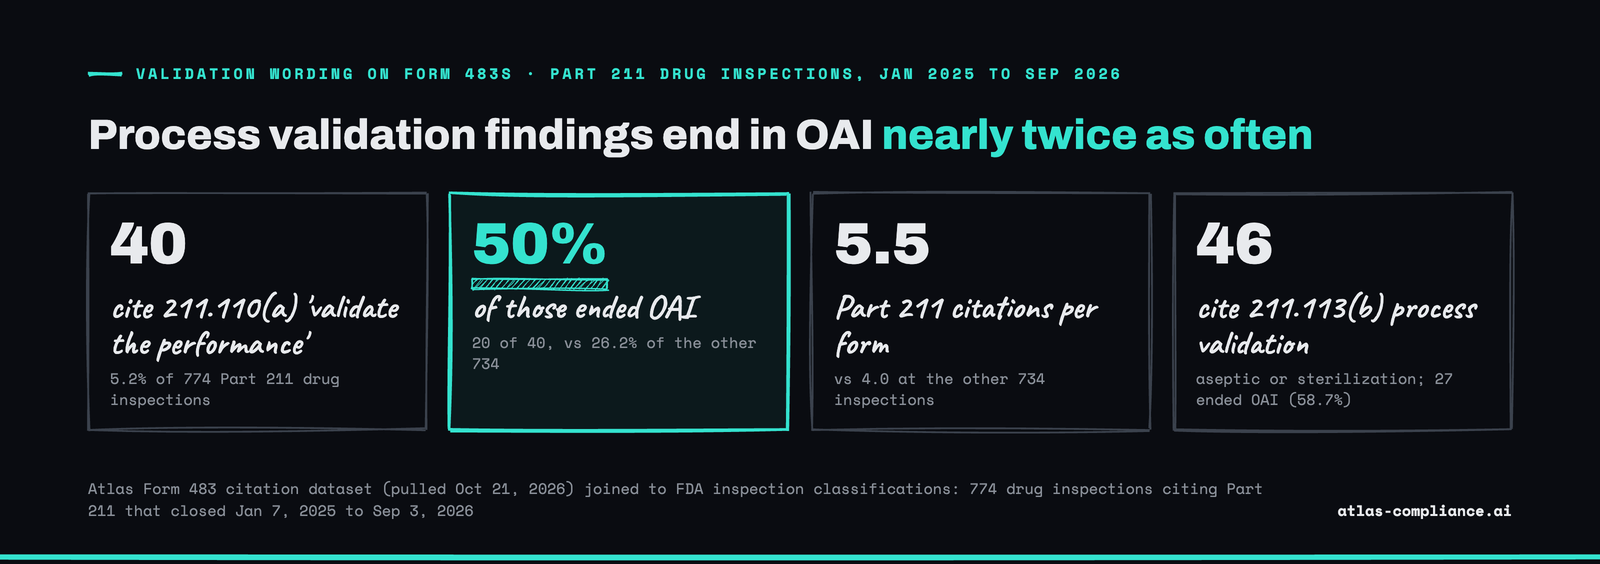
- 40 of 774 Part 211 drug inspections (5.2%) carried the 21 CFR 211.110(a) citation that control procedures were not established to "validate the performance" of manufacturing processes.
- 20 of those 40 inspections (50%) ended Official Action Indicated (OAI), against 192 of the other 734 (26.2%).
What is process performance qualification (PPQ)?
Process performance qualification is the point in the product lifecycle where a manufacturer proves the commercial process works as designed, at commercial scale, before selling product. FDA's 2011 guidance describes PPQ as combining the qualified facility, utilities and equipment and the trained personnel with the commercial manufacturing process, control procedures and components to produce commercial batches.
Stage 1 designs the process, Stage 2 qualifies it and Stage 3 keeps it in control; our process validation guide covers all three.
The guidance states that process validation, including process qualification, is legally enforceable under section 501(a)(2)(B) of the Federal Food, Drug, and Cosmetic Act. 21 CFR 211.110(a) supplies the wording investigators use: control procedures "shall be established to monitor the output and to validate the performance of those manufacturing processes that may be responsible for causing variability."
What must a PPQ protocol contain?
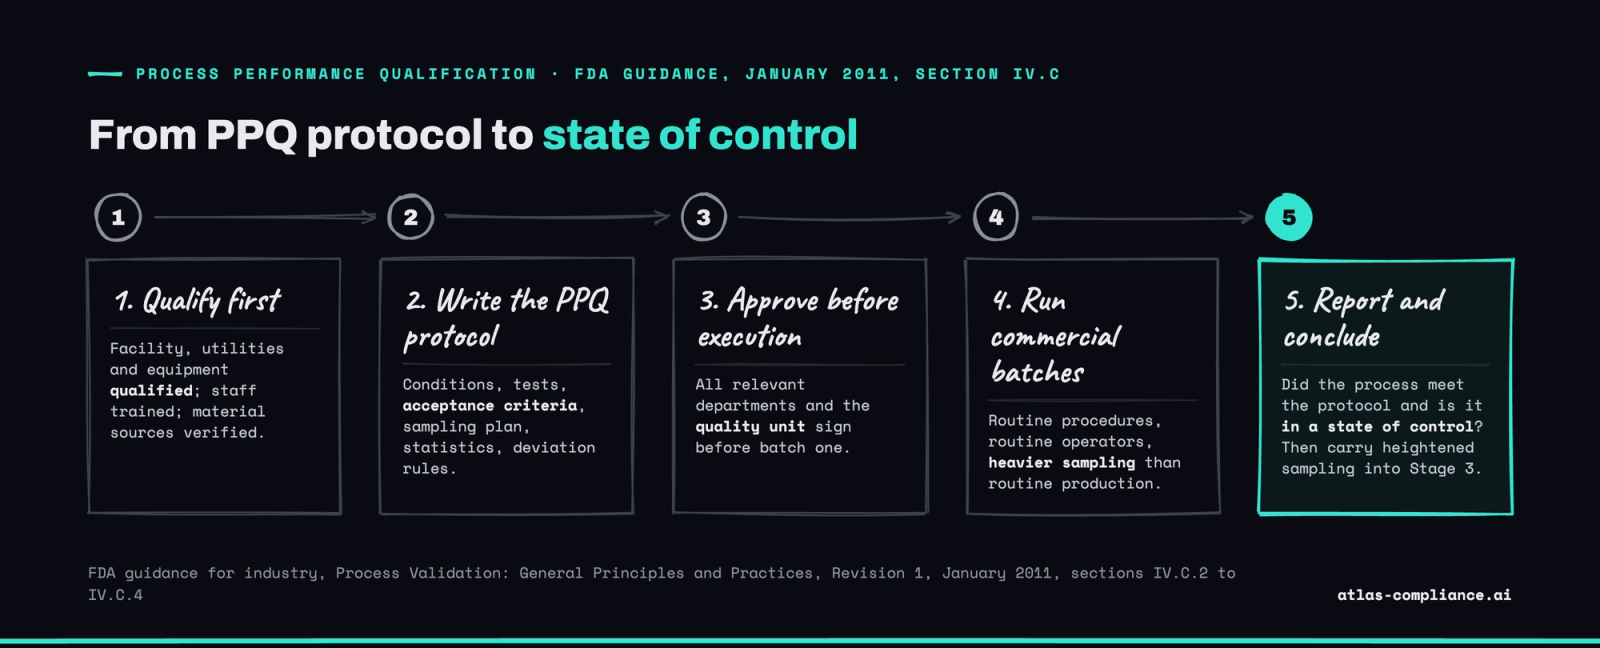
A PPQ protocol must specify the manufacturing conditions, controls, tests and expected outcomes before the first batch is made. FDA's 2011 guidance, section IV.C.3, recommends that the protocol cover these elements:
- Manufacturing conditions: operating parameters, processing limits and component (raw material) inputs.
- The data to collect, and when and how your team will evaluate it.
- In-process, release and characterization tests, with acceptance criteria for each significant processing step.
- A sampling plan with sampling points, number of samples and sampling frequency for each unit operation and attribute.
- Criteria and process performance indicators, including the statistical methods for intra-batch and inter-batch variability and the rules for deviations and nonconforming data.
- Facility, utility and equipment qualification, personnel training and verification of material sources, if not already done.
- The validation status of the analytical methods used to measure the process, in-process materials and product.
- Review and approval by the relevant departments and the quality unit.
Firms that also supply the EU add the items in Annex 15, paragraph 5.22, among them the master batch record reference, the critical quality attributes (CQAs) and the critical process parameters (CPPs) with their limits.
For execution, the guidance says no batch should start until every relevant department, including the quality unit, has approved the protocol. The operators who run each step in routine production make the PPQ lots under normal conditions, following the commercial procedures required by 211.100(b) and 211.110(a). Any departure from the protocol needs justification and approval from those same groups before anyone acts on it.
The guidance then asks for a timely PPQ report that cross-references the protocol, analyzes the data as the protocol specified, discusses every deviation and aberrant result, and states a clear conclusion on whether the process met the protocol and is in a state of control. If the answer is no, the report says what has to happen before the team can reach that conclusion.
How many PPQ batches are required?
The number of PPQ batches required is whatever the manufacturer can justify from its product and process knowledge, because FDA's 2011 guidance does not set one. The guidance asks for sampling that gives statistical confidence within and between batches. EU GMP Annex 15 treats three consecutive batches as a generally acceptable minimum, subject to a risk-based justification.
| Question | FDA guidance (January 2011) | EU GMP Annex 15 (2015) |
|---|---|---|
| Fixed batch number | None stated | None fixed; 3 consecutive batches "generally considered acceptable" (5.20) |
| Basis for the number | Sound science, product and process understanding, demonstrable control | Quality risk management; normal range of variation and trends (5.19) |
| Who justifies it | The manufacturer, in the protocol | "Each manufacturer must determine and justify" (5.19) |
| Fewer or more than three | Not addressed as a number | Alternative number may be justified by standard methods or similar products at the facility (5.20) |
| After the initial batches | Heightened sampling continues into Stage 3 | Initial three batches may need supplementing with ongoing process verification data (5.20) |
The guidance names the inputs that move the number. Your team has to consider the effects of scale, but need not explore the full operating range at commercial scale where process design data already give assurance. Credible experience with similar products helps, and FDA recommends statistical metrics wherever feasible. A new dosage form with little scale-up data has to buy its confidence with more batches or more samples per batch; a familiar process can lean on its history.
A firm selling into both markets has to meet Annex 15, so three consecutive batches becomes its working floor unless the protocol makes the statistical case for another number. An investigator reviewing your PPQ will ask to see that reasoning.
How do you set PPQ sampling and acceptance criteria?
PPQ sampling and acceptance criteria come from the protocol's statistical plan: sample at every unit operation and attribute that can vary, take more samples than routine production does, and set the confidence level for each attribute by risk analysis. Testing should confirm uniform quality throughout each batch, and your team cannot exclude PPQ data without a documented, science-based justification.
Acceptance criteria for in-process tests connect to the regulation. 21 CFR 211.110(b) requires in-process specifications consistent with final product specifications, derived from process average and variability estimates where possible and set by suitable statistical procedures where appropriate. 211.110(a) names examples such as weight variation, adequacy of mixing, dissolution and bioburden.
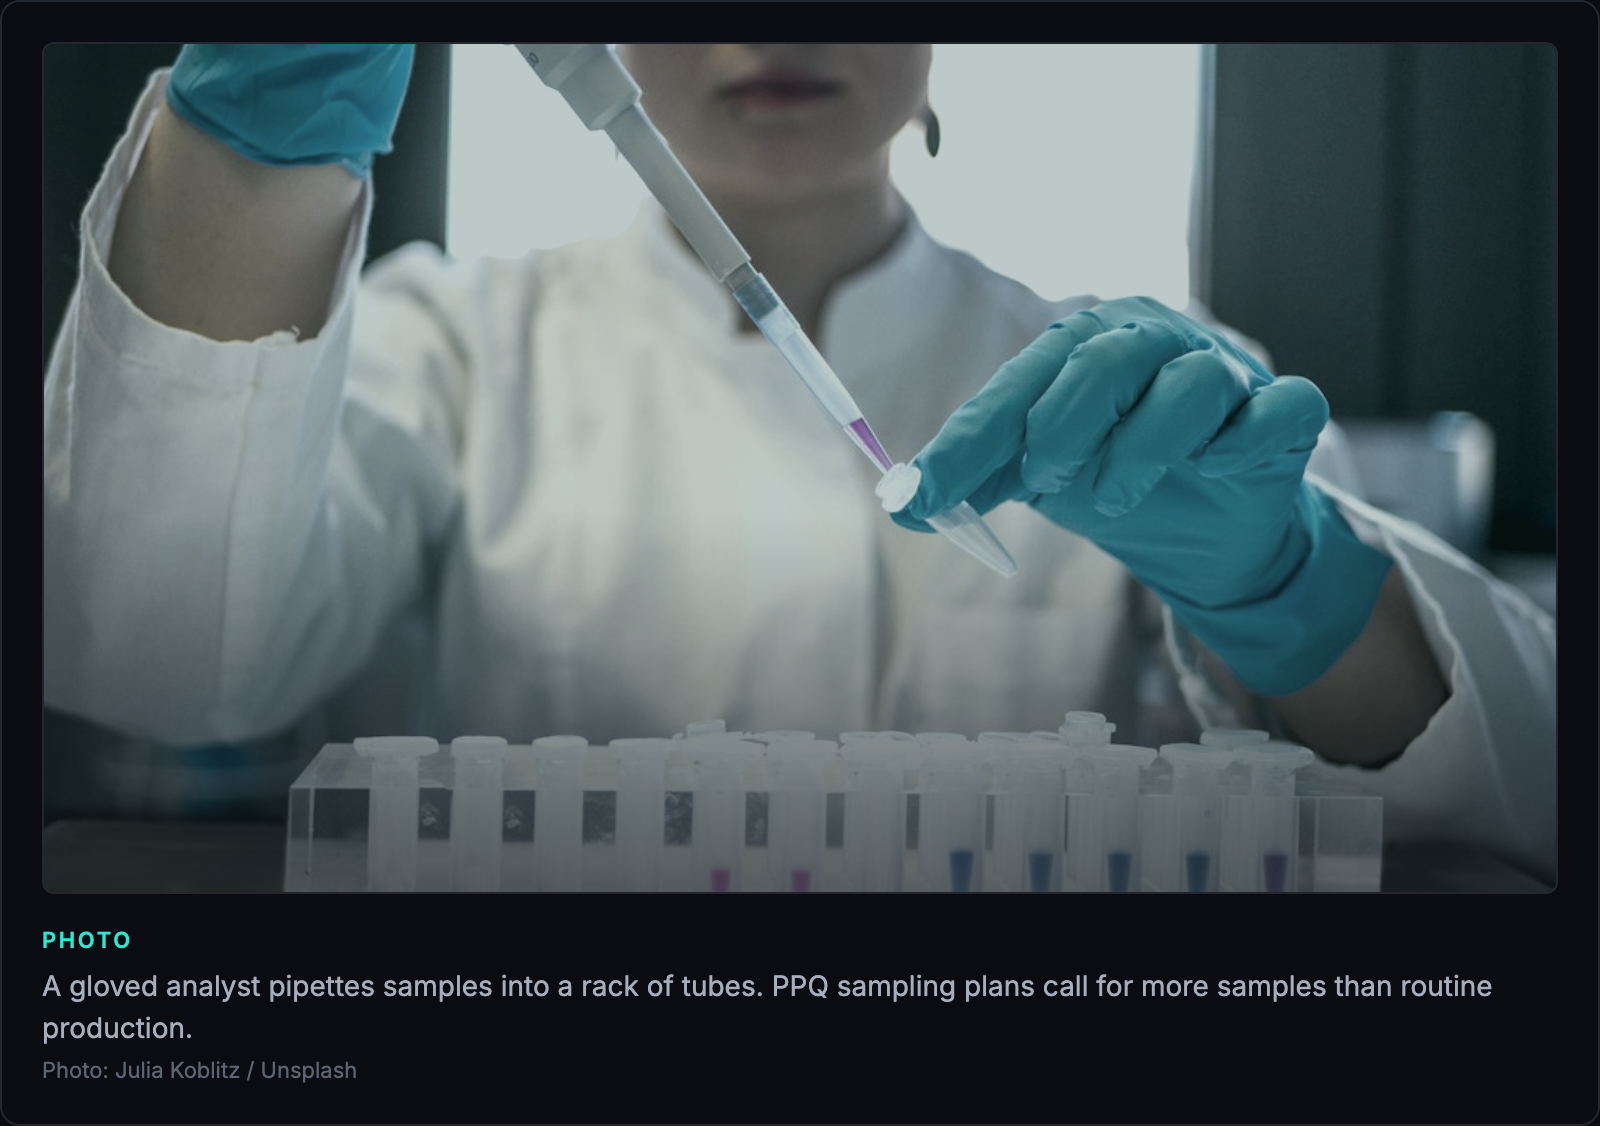
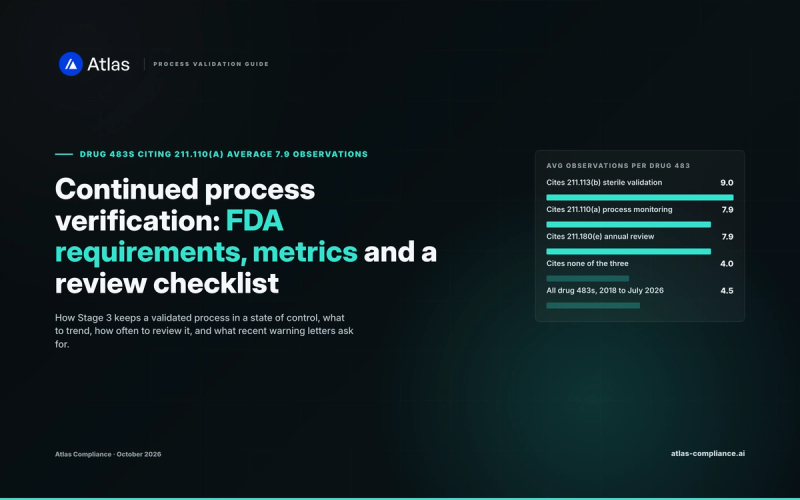
The guidance recommends keeping PPQ-level monitoring into Stage 3 until there are enough data to estimate variability, then setting routine sampling from those estimates. Our continued process verification guide covers that handover.
PPQ vs IQ, OQ and PQ: what is the difference?
PPQ qualifies the whole commercial process, while IQ, OQ and PQ qualify the individual equipment and utilities that process runs on. Both sit in Stage 2 of FDA's 2011 guidance. Equipment and utility qualification is the first element and must come before PPQ; PPQ is the second element and must come before commercial distribution.
| Aspect | Equipment qualification (IQ, OQ, PQ) | Process performance qualification |
|---|---|---|
| Scope | One system: a granulator, autoclave, HVAC or water system | The full commercial process, all unit operations |
| Question answered | Is it installed to specification and does it operate across all anticipated ranges? | Does the process reproduce quality product at commercial scale? |
| Conditions | Load comparable to routine production, including interventions, stoppage and start-up | Normal conditions, routine operators, commercial components and batch size |
| Approval | Quality control unit approves the plan and report (211.22) | All relevant departments and the quality unit approve the protocol before execution |
| Comes before | PPQ | Commercial distribution |
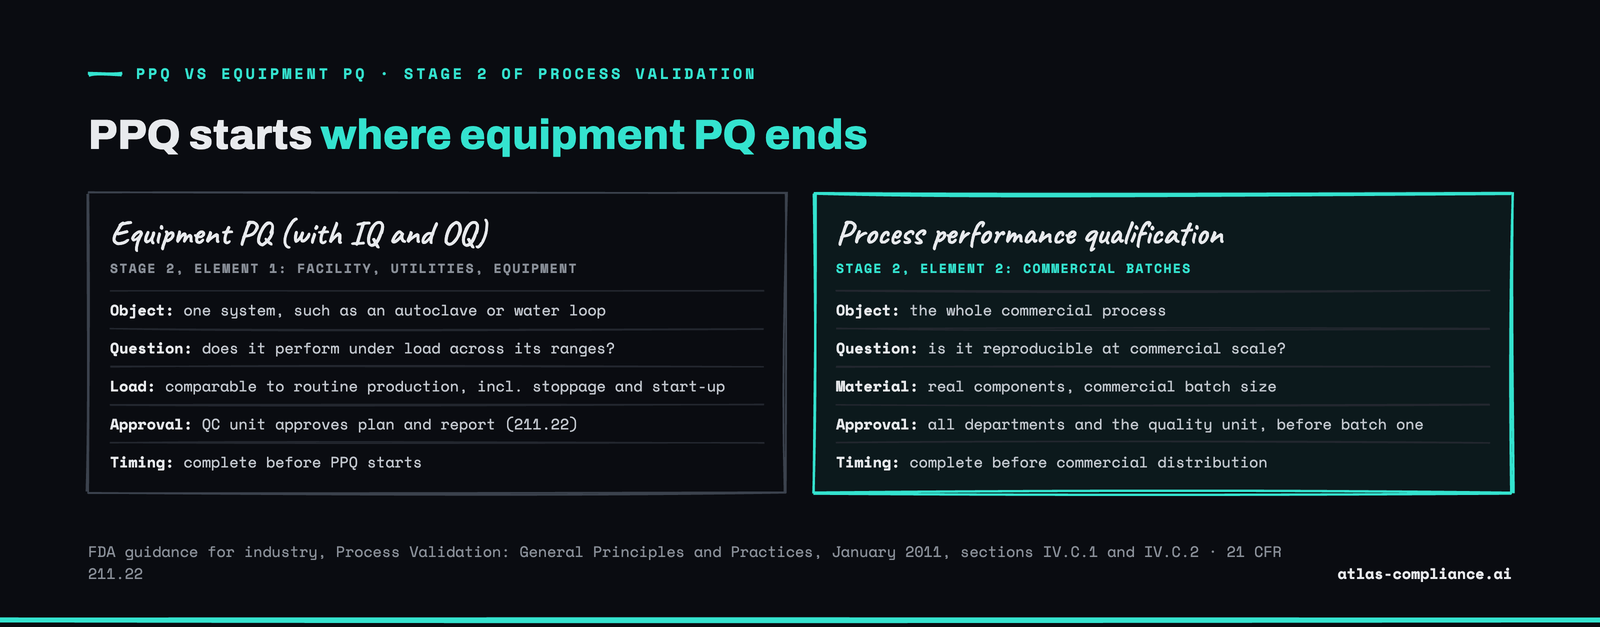
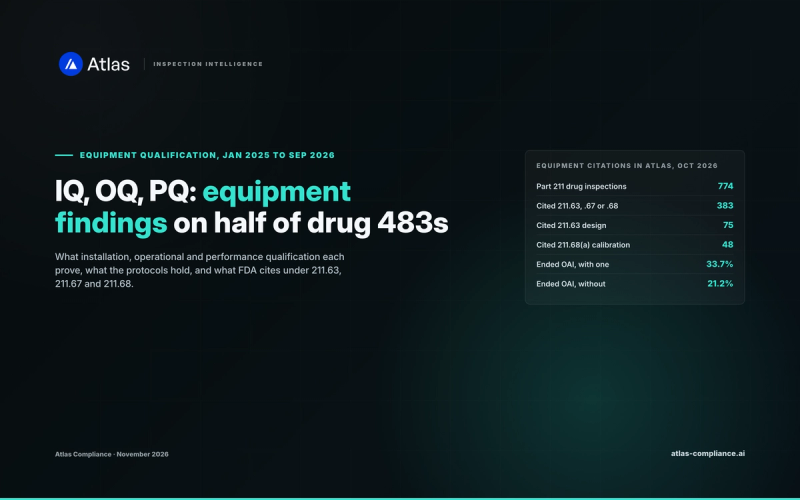
The abbreviations collide: in equipment work PQ means performance qualification of one system, while the 2011 guidance uses PQ for the whole process qualification stage. Our IQ, OQ and PQ guide covers what each equipment protocol holds.
Can PPQ batches be sold?
PPQ batches can be sold if they meet their release criteria, because the 2011 guidance states that products made during Stage 2, if acceptable, can be released for distribution. The guidance adds that in most cases the PPQ study should be completed successfully before commercial distribution starts.
Releasing a PPQ batch before the protocol is fully executed is concurrent release, and FDA expects firms to use it rarely: for orphan or minor species drugs with limited demand, short half-life products such as positron emission tomography drugs, and medically necessary drugs made in coordination with FDA during a shortage. The protocol has to state the rationale, and each lot still has to meet every CGMP requirement and the protocol's lot release criteria. Annex 15, paragraph 5.14, adds that release of validation batches to the market should be pre-defined.
What does FDA cite when PPQ evidence is missing?
When a facility cannot show its process performance was validated, FDA investigators cite 21 CFR 211.110(a) with the wording "Control procedures are not established which monitor the output and validate the performance of those manufacturing processes." In the Atlas Form 483 citation dataset (pulled October 21, 2026), 40 of 774 Part 211 drug inspections (5.2%) carried that wording.
The 40 inspections split 32 in 2025 (5.7% of 562 Part 211 drug inspections) and 8 from January to September 2026 (3.8% of 212). Thirty-four were at US facilities and six at foreign ones. They ended worse than the rest: 20 of 40 were classified OAI, against 26.2% of the other 734 inspections, and the forms averaged 5.5 Part 211 citations against 4.0.
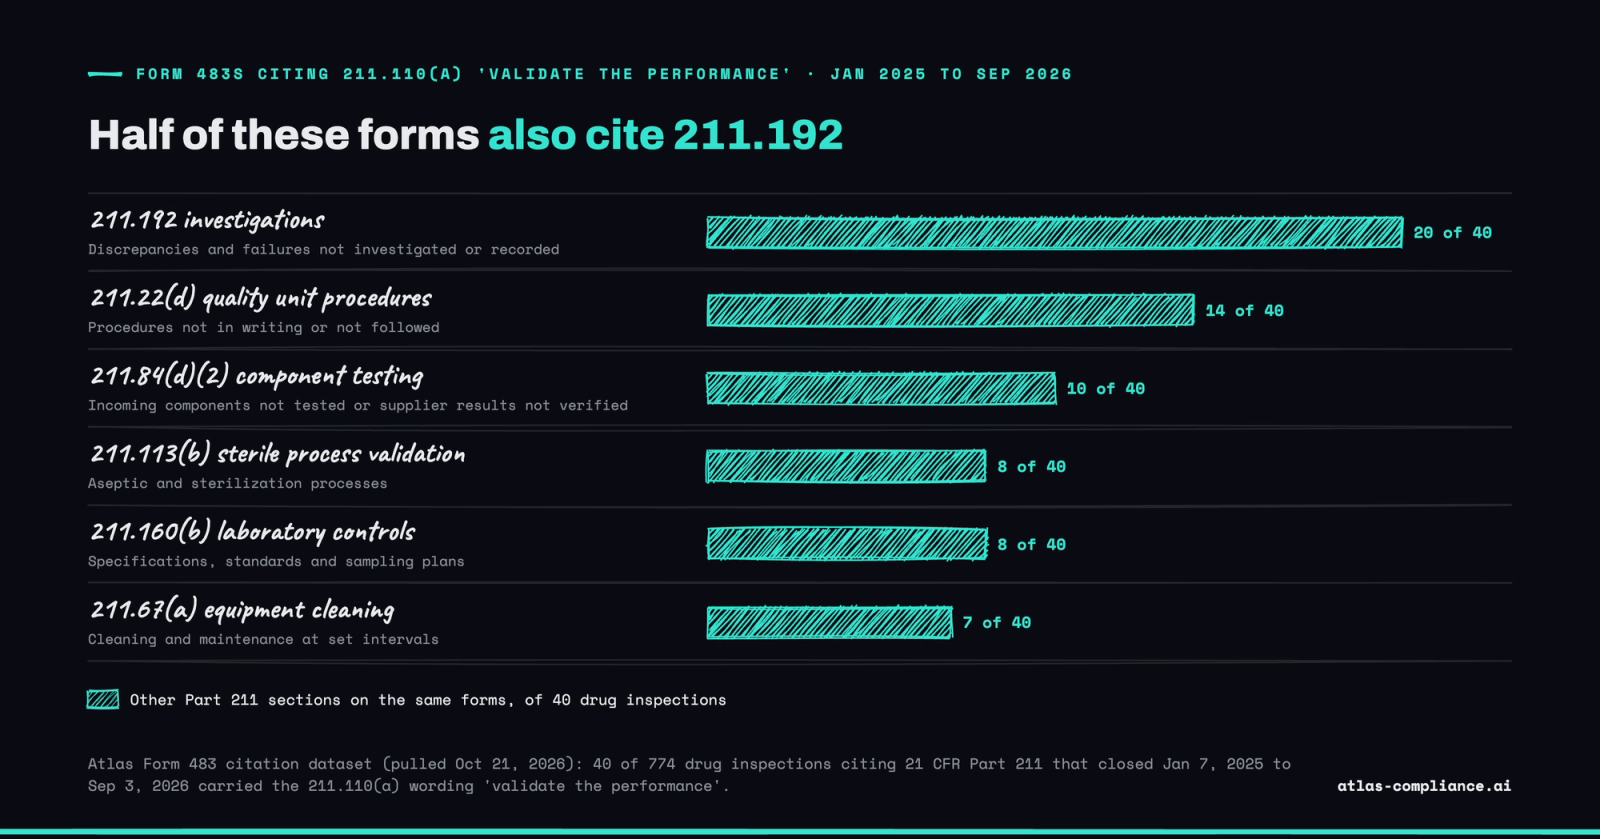
None of the 40 inspections cited the validation finding alone; each carried at least one other Part 211 section. Half of the 40 also cited 21 CFR 211.192, the investigations section covered in our 211.192 guide, and 14 cited 211.22(d) on quality unit procedures.
Five forms from the FDA Form 483 database show the range:
| Form 483 | Inspection closed | Classification | Other Part 211 sections on the form |
|---|---|---|---|
| K.C. Pharmaceuticals, Pomona, US | Feb 13, 2026 | OAI | 211.113(b), 211.192, 211.42(c)(10), 211.68(b) and 8 more |
| Grand River Aseptic Manufacturing, Caledonia, US | Nov 4, 2025 | VAI | 211.22(d) |
| Frontida Biopharm, Aurora, US | Oct 24, 2025 | VAI | 211.100(b), 211.22(a), 211.22(d) |
| Evaric Pharmaceuticals, Hauppauge, US | May 16, 2025 | VAI | 211.42(c), 211.67(a) |
| Epic Laboratories, Tampa, US | Apr 2, 2025 | OAI | 211.113(a), 211.160(b), 211.166(b), 211.192, 211.22(d), 211.67(a) |
The K.C. Pharmaceuticals form is the largest of the five: 13 observations, with the process validation citation sitting next to a 211.113(b) finding that aseptic process validation was inadequate. The K.C. Pharmaceuticals Pomona facility has 16 FDA inspections on record, 11 ending with a Form 483. Grand River's form shows the other end: two observations, the validation finding and a quality unit procedure citation, and a VAI outcome.
Sterile manufacturers face a second validation line. 21 CFR 211.113(b) appeared with validation wording at 46 of the 774 inspections, 27 of which (58.7%) ended OAI. Foreign facilities drew it at 23 of 142 inspections (16.2%), against 23 of 632 US inspections (3.6%).
Each investigator on these forms has an inspection history in Atlas. Before a pre-approval or surveillance inspection, you can check which validation sections your likely investigator has cited at peer facilities.
 Atlas
See what the FDA knows about your next investigator.
30 minutes with the founder. No pitch deck.
When does PPQ need to be repeated?
PPQ needs to be repeated, in full or in part, when a change or a Stage 3 signal means the original PPQ data no longer represent the commercial process. FDA's 2011 guidance says that, depending on how a proposed change might affect product quality, additional process design and process qualification activities could be warranted.
FDA and EU texts point to four triggers:
- Planned changes to operating conditions, process controls, components or in-process material characteristics. The guidance requires a description, rationale, implementation plan and quality unit approval before implementation under 211.100, and footnote 19 points to 21 CFR 314.70 and 601.12 for changes that need FDA notification.
- A failed PPQ. The guidance says additional design studies and qualification may be necessary if Stage 2 does not show reproducible performance at commercial scale.
- Requalification of equipment and facilities. The guidance asks firms to assess qualification data periodically and decide whether re-qualification is needed and how much.
- Changes to starting materials, equipment, premises, batch size or method of production for EU supply. Annex 15, paragraphs 11.2 and 11.4, calls for quality risk management to plan any process validation, verification or requalification such a change needs.
Annex 15, paragraph 5.27, also lets a firm use a hybrid of traditional and continuous verification for validation after a change, even for a product first validated with traditional batches. Our post on change control in project management covers how to route those decisions.
If your next PPQ follows a change, check its protocol against the 211.110(a) wording above before the inspection. The digest below sends new process validation findings as Atlas indexes them.
AtlasFDA Inspection Intelligence Digest
We track every 483, EIR, and warning letter the moment it happens. Get the signals before they hit the FDA website.
Frequently asked questions
PPQ is one part of process validation. FDA's 2011 guidance defines process validation as three stages: process design, process qualification and continued process verification. Process qualification, Stage 2, has two elements, and PPQ is the second, after facility, utility and equipment qualification. A completed PPQ shows the process can run at commercial scale. The firm still has to keep it in a state of control through Stage 3 monitoring.
Written by
Bhavish Agarwal
Founder, Atlas
Bhavish Agarwal is the founder of Atlas and previously worked at Google and Microsoft. He has spent the past several years building regulatory data infrastructure for the pharmaceutical industry.