
Continued process verification (CPV) is Stage 3 of FDA's process validation lifecycle: the ongoing collection and analysis of production data that shows a commercial process stays in a state of control after process performance qualification (PPQ). It runs for as long as you make the product, and it follows the process through supplier changes, equipment wear and routine adjustments.
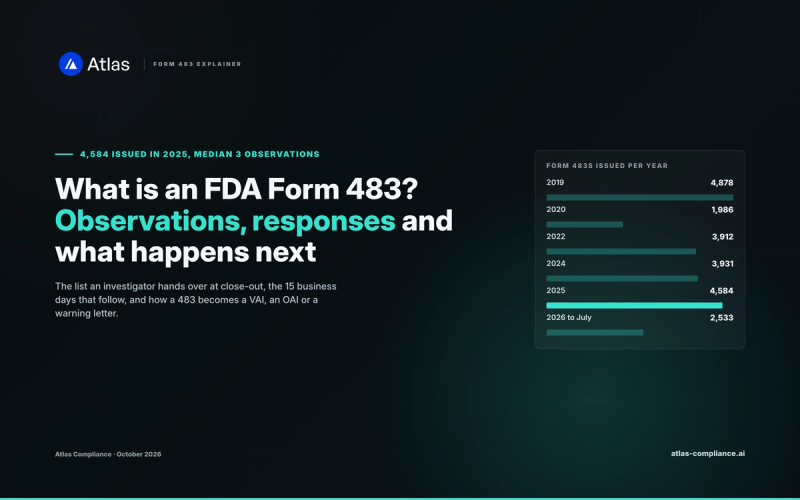
On a Form 483, investigators cite the regulations CPV exists to satisfy. Across 5,177 human-drug Form 483s issued since 2018 in the Atlas directory, 306 cite 21 CFR 211.110(a), the requirement to monitor and validate the performance of manufacturing processes. Those 483s average 7.9 observations. Drug 483s that cite none of the process-validation sections average 4.0.
What FDA requires for continued process verification
The 2011 guidance, Process Validation: General Principles and Practices, defines the three stages: process design, process qualification, and continued process verification. The guidance is a recommendation; the regulations it rests on are binding.
| Requirement | Source | What it means for CPV |
|---|---|---|
| Written production and process control procedures designed to assure quality | 21 CFR 211.100(a) | The CPV plan, its sampling and its review steps are written procedures, approved by the quality unit |
| Control procedures to monitor output and validate the performance of processes that may cause variability | 21 CFR 211.110(a) | You trend in-process and finished-product data and look for variability as well as failures |
| Records that allow at least an annual evaluation of quality standards for each product | 21 CFR 211.180(e) | The annual product review draws on CPV data. It does not replace it |
| Review of production records, and investigation of any unexplained discrepancy | 21 CFR 211.192 | A CPV signal that points at a discrepancy needs an investigation on record |
| Ongoing process verification to keep a state of control through the product lifecycle | EU GMP Annex 15, section 5 | The European name for the same stage. Global sites run one program for both |
| A process performance and product quality monitoring system | ICH Q10 | The quality system that owns CPV and feeds its outcomes into CAPA and change management |
The guidance also recommends three practices. Keep sampling at the level used during PPQ until you have enough data to estimate variability with confidence. Have a statistician, or someone trained in statistical process control, design the data collection plan. Look at both intra-batch and inter-batch variation.
How often do drug 483s cite process monitoring?
We counted three citations across the human-drug Form 483s in the Atlas Form 483 database, using FDA's own citation names (data through July 2026):
| Year | Human-drug 483s | Cite 211.110(a), monitor and validate performance | Share |
|---|---|---|---|
| 2018 | 809 | 56 | 6.9% |
| 2019 | 859 | 68 | 7.9% |
| 2020 | 261 | 12 | 4.6% |
| 2021 | 388 | 25 | 6.4% |
| 2022 | 548 | 31 | 5.7% |
| 2023 | 601 | 47 | 7.8% |
| 2024 | 633 | 25 | 3.9% |
| 2025 | 755 | 35 | 4.6% |
| 2026 (to July) | 323 | 7 | 2.2% |
Over the same period, 112 drug 483s cite 21 CFR 211.180(e), items to cover on annual reviews, and 207 cite 21 CFR 211.113(b), validation lacking for sterile drug products.
The share citing 211.110(a) fell after 2023, but the 483s that do cite it are heavy. A quarter of them carry ten or more observations, against 9% of all drug 483s. They also cite the neighboring controls more often: investigations of discrepancies (211.192) on 29% of them against 16% of other drug 483s, laboratory controls (211.160) on 31% against 13%, and procedures not followed (211.100(b)) on 39% against 24%.
Three recent examples:
- K-C Pharmaceuticals, February 13, 2026: 13 observations, including 211.110(a).
- Teva Women's Health, March 20, 2026: 2 observations, one of them 211.110(a).
- Hospira, January 17, 2025: 3 observations, including 211.110(a).
What a CPV program contains
A CPV program is a written plan for each product. At minimum it names:
- The data. Critical quality attributes, critical process parameters, in-process results, yields, and the raw-material attributes that affect them.
- The split. How you separate the data by line, equipment train, raw-material supplier and lot, shift or campaign. A pooled view hides local problems.
- The statistics. Control charts with defined run rules, capability indices, and the batch count at which you recalculate limits.
- The cadence. How often each data set is reviewed, and by whom.
- The triggers. Which signals open a review, and which outcomes the reviewer can choose.
- The link to change control. Every approved change to the process names the batches and data that will confirm it preserved performance.
Which metrics to trend
| Metric | What it shows | Example trigger |
|---|---|---|
| Individuals or X-bar control chart per CQA | Shifts and trends inside specification | A run rule fires: eight consecutive points on one side of the mean, or six in a row rising |
| Range or standard deviation chart | Variability widening while the mean holds | The range chart breaks its control limit |
| Process capability (Cpk, Ppk) | Margin between the process and the specification | Ppk drops below the plan's target, for example 1.33 |
| Out-of-trend results | Results that are in specification but unusual for this process | Any confirmed out-of-trend result |
| Deviation rate by process step | Repeated small failures in one place | Two or more minor deviations on the same step within a review period |
| Yield and reject rate | Losses the specification does not see | A sustained drop against the PPQ baseline |
| Post-change batches | Whether a change behaved as predicted | Any post-change batch outside the PPQ range |
Read the control chart before the capability index. Cpk and Ppk assume the process is in statistical control.
How often to review CPV data
FDA leaves the review interval to you. Set it by risk and production rate. We suggest this as a starting point:
| Data | Suggested review | Why |
|---|---|---|
| Critical quality attributes on high-risk or sterile products | Every batch, trend reviewed monthly | A drift can reach a patient within a few batches |
| Other CQAs and CPPs | Monthly or quarterly | Frequent enough to catch a trend before the annual review |
| New products, and the batches after a change | Every batch until the planned batch count is met | Variability estimates are still forming |
| Low-volume products | Per campaign | Too few batches for a monthly chart to mean anything |
| Whole-product summary | Annually, in the annual product review (211.180(e)) | The required yearly look, built on the reviews above |
If your team first discusses a drift at the annual product review, that drift has had a year of batches to grow.
Escalation rules: every CPV signal ends in a decision
For each signal, the reviewer records one of five outcomes:
- No action, with a scientific rationale.
- Enhanced monitoring for a defined number of batches.
- Investigation or CAPA, through the normal quality system.
- A change to the sampling plan or the control strategy, through change control.
- Targeted requalification or revalidation of the affected step.
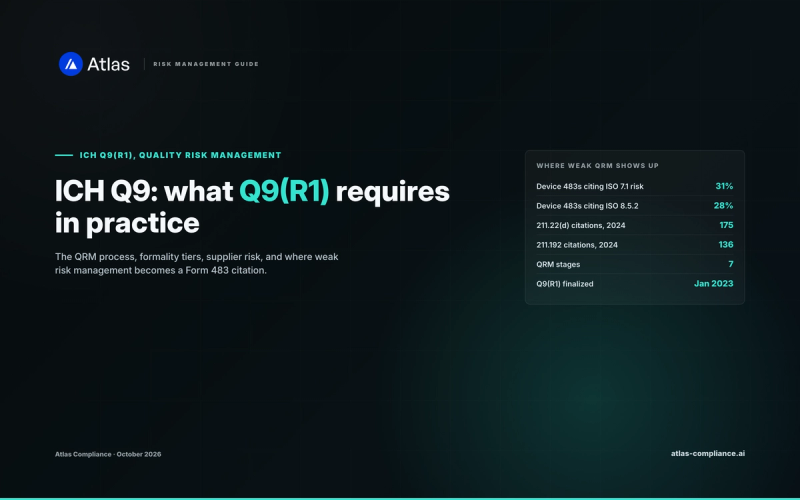
Name an owner and a date for each outcome. The record should let another reviewer, including an FDA investigator, follow the data to the decision. Rank signals with the same risk logic you use elsewhere in the quality system; our guide to ICH Q9 quality risk management covers the method.
What FDA asked for in 2026 warning letters
In two recent letters, FDA spelled out the evidence it expects when process validation fails.
On May 29, 2026, FDA's warning letter to Macau-Union Pharmaceutical asked the firm to describe its program for PPQ and for ongoing monitoring of both intra-batch and inter-batch variation to ensure a continuing state of control. It also asked for lifecycle oversight of equipment, cleaning and changes.
On April 9, 2026, FDA's letter to Medical Products Laboratories in Philadelphia asked for PPQ, procedures, risk assessments for products made with inadequately validated processes, and ongoing monitoring that can show a continuing state of control.
In both letters, FDA asked the firm to show how it knows the process is in control today. A CPV program is the record that answers that question. Our guide to FDA Form 483s explains how an observation becomes a letter like these.
 Atlas
See what the FDA knows about your next investigator.
30 minutes with the founder. No pitch deck.
CPV vs. continuous process verification vs. annual product review
Teams often use these four terms interchangeably. Each means something different:
| Term | What it is | When it applies |
|---|---|---|
| Continued process verification (CPV) | FDA Stage 3: ongoing monitoring of a validated commercial process | After PPQ, for the life of the product |
| Ongoing process verification (OPV) | The EU GMP Annex 15 name for the same stage | Products made for the EU market |
| Continuous process verification | An alternative to traditional validation batches, using real-time monitoring of the process (ICH Q8, Annex 15) | In place of a conventional PPQ, built on process analytical technology |
| Annual product review (APR) | A yearly review of records for each product, required by 21 CFR 211.180(e) | Every year, for every product |
CPV review checklist
Use this at each scheduled review:
- Confirm every CQA and CPP in the plan has current data, with no gaps.
- Split the data by line, equipment, material supplier and lot, as the plan requires.
- Update the control charts and apply the run rules.
- Recalculate capability indices where the plan calls for it, using in-control data.
- List each out-of-trend result with its disposition.
- Map the period's deviations to process steps and flag repeats.
- List the changes approved in the period, with their post-change batch data.
- Give each signal one of the five outcomes, an owner and a date.
- Close the open actions from the last review, or carry them forward with a reason.
- Get the quality unit's signature on the review.
For the CFR sections that come up most often on drug 483s, see our analysis of the most prevalent CFR citations.
AtlasFDA Inspection Intelligence Digest
We track every 483, EIR, and warning letter the moment it happens. Get the signals before they hit the FDA website.
Frequently asked questions
The term comes from FDA guidance, which is a recommendation. The regulations behind it are binding: 21 CFR 211.110(a) requires control procedures that monitor and validate process performance, and 21 CFR 211.180(e) requires records that support at least an annual quality evaluation. Investigators cite a firm without ongoing process monitoring under those sections, whatever the firm calls its program.
Written by
Bhavish Agarwal
Founder, Atlas
Bhavish Agarwal is the founder of Atlas and previously worked at Google and Microsoft. He has spent the past several years building regulatory data infrastructure for the pharmaceutical industry.